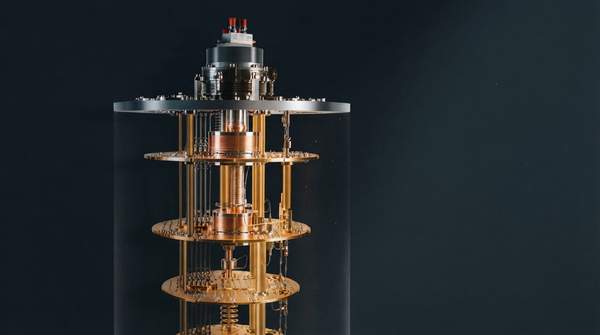
Quantum computers hold a structural advantage for molecular simulation that classical machines cannot copy: they run on the same quantum-mechanical rules that govern chemistry itself. A 2026 review in Frontiers in Drug Discovery, authored by Harshraj Gadbail, Rajendra Rewatkar, Nishant Jumde, Mrinal Manker, Pratik Nagre and Sujal Zade, frames the field's open question precisely — under what conditions do quantum methods offer a tangible advantage over the best available classical approaches — and answers it task by task. The short version: today's quantum algorithms such as the Variational Quantum Eigensolver (VQE) already match classical accuracy on very small molecules, while future algorithms like Quantum Phase Estimation (QPE) are the ones positioned to extend high-accuracy energy calculations to drug-sized molecules that the classical gold standard, CCSD(T), cannot reach.
The Limits of Classical Simulation in Drug and Materials Science
The core bottleneck is electron correlation. Computer-aided drug design has transformed how researchers navigate chemical space and predict ligand–target interactions, but as the Frontiers review explains, classical implementations rely on mechanical force-field approximations that fail to capture polarisation, charge transfer, and electron correlation — the effects central to molecular recognition and reactivity. Quantum-mechanical treatments that describe these phenomena correctly scale exponentially with system size on classical hardware, rendering them impractical for drug-relevant biomolecules.
The scaling numbers make that wall concrete. According to the Frontiers comparison, classical density functional theory (DFT) with hybrid functionals achieves roughly 1–3 kcal/mol accuracy for small molecules but scales as O(N³), becoming impractical beyond about 100–200 atoms. The "gold standard" coupled cluster method, CCSD(T), reaches sub-kcal/mol accuracy but scales as O(N⁷), restricting its use to roughly 20–30 atoms. The trade-off is stark: chemists can have either broad applicability with approximate accuracy, or exquisite accuracy on systems small enough to fit in a fragment of a protein.
That leaves a structural gap. The systems that matter most for drug discovery and materials science — metalloprotein active sites, reactive intermediates, nanomaterials — sit well beyond both limits while demanding exactly the accuracy these methods sacrifice at scale. Metal-containing proteins, for instance, are precisely where electron correlation, charge transfer and polarisation dominate the physics, which is why the standard classical toolkit degrades fastest on the hardest targets. Any technology that could represent correlated electrons natively, rather than approximating them, would attack the gap at its widest point.
Quantum Advantage: Superposition and Entanglement for Molecular Simulation
Why should different hardware help? Because the problem and the machine share a substrate. As St. Jude researcher Arpad Gorgulla put it in a 2025 piece by Alex Generous, chemistry is the interaction of molecules at the subatomic level — in particular, the electron level — and those interactions are best described by quantum mechanics. Quantum computers are designed on those same principles, which makes them better suited for analyzing molecular interactions than classical machines.
The mechanics translate directly. Classical computers use bits that are either zero or one, "like a light switch," performing operations in stepwise arithmetic that becomes very time-consuming as calculations grow complex, Gorgulla explained. Qubits instead exploit superposition and entanglement — the two principles Drug Discovery World's Diana Turner identifies as the engine of the quantum advantage. Superposition lets a quantum state represent many configurations of a molecular wavefunction simultaneously, which maps naturally onto exploring chemical space rather than enumerating it one configuration at a time. Entanglement lets correlated qubits encode the linked behavior of interacting electrons — precisely what force fields and approximate classical methods miss.
The payoff, in Gorgulla's framing, is that quantum computers can in theory be exponentially faster for the thermodynamic and quantum-mechanical calculations required to simulate a protein and all molecules that could potentially bind to it. The important caveat is the word "theory": the advantage follows from shared physics, not from any demonstrated benchmark at drug-relevant scale. Classical computers can, in principle, perform the same simulations; the computational cost is what becomes impractical, especially with the highly accurate quantum-mechanical methods that matter when modeling molecular interactions with high precision.
Quantum Algorithms for Drug Discovery and Materials Science
Matching algorithm to task is where the field becomes practical. VQE is a hybrid quantum-classical algorithm designed for near-term quantum devices: a quantum processor prepares trial molecular states, and a classical optimizer iterates toward the lowest energy — the ground state. As a review in the National Library of Medicine's archive puts it, VQE is particularly useful for simulating the electronic structure of molecules, leveraging quantum mechanics to find the ground state, a critical task for predicting how molecules will interact in biological systems.
QPE is the more demanding sibling. It targets the same ground-state energies but requires the fault-tolerant quantum computers of the future, and in exchange delivers CCSD(T)-grade accuracy on molecules far larger than 30 atoms — the Frontiers table places this capability after 2030. The division of labor is therefore temporal: VQE is the algorithm for today's noisy hardware on toy systems, while QPE is the algorithm whose payoff arrives once error-corrected machines exist.
Beyond energies, the same review notes that Grover's search algorithm can scan vast chemical databases to identify candidates matching specific molecular criteria, with a quadratic speedup over classical search in theory. Quantum machine learning models such as quantum support vector machines and quantum neural networks round out the toolkit, though the Frontiers assessment is blunt: they have not yet beaten classical AI on property prediction, and no clear advantage has been demonstrated.
Quantum vs. Classical: A Comparative View of Molecular Simulation
Juxtaposing the scaling and accuracy limits of each classical method against its quantum counterpart shows a consistent pattern: quantum methods do not replace classical chemistry wholesale — they take over where classical accuracy collapses. The table below consolidates the Frontiers comparison across the primary computational tasks in drug discovery, pairing each classical baseline with the quantum alternative and its readiness.
| Computational Task | Classical Method | Quantum Method | Limitations & Target Scale |
|---|---|---|---|
| Calculating molecule energy (small molecules) | DFT (standard method) — good, within 1–3 energy units | VQE — works on today's quantum computers, similar accuracy, slightly larger molecules | DFT slows beyond ~100 atoms; VQE limited to toy systems (<30 atoms); available now at limited scale |
| Calculating molecule energy (large, complex molecules) | CCSD(T) (most accurate classical method) — excellent accuracy | QPE — same high accuracy for much larger drug molecules | CCSD(T) only works up to ~30 atoms; QPE needs future quantum computers (after 2030) |
| Predicting how strongly a drug binds to its target | FEP+ (industry standard on GPU computers) — good, within ~1 energy unit for similar drug series | Quantum-assisted QM/MM for the binding site — more accurate for difficult metal-protein targets | FEP+ struggles with metal-containing proteins and reactive drugs; quantum approach still future research |
| Screening millions of drug candidates quickly | Docking software (Glide, AutoDock) — rough estimates only | Grover's search algorithm — faster database search (quadratic speedup in theory) | Docking misses many good candidates; Grover needs future quantum computers (after 2030) |
| Predicting drug properties from molecular structure | Deep learning / AI models (graph neural networks) — excellent, state of the art | Quantum machine learning (QSVM, QNN) | Classical AI needs large datasets and is black-box; quantum ML not yet proven to beat it — research stage |
Read row by row, the table yields a clear evidence-derived result: for tasks requiring high-fidelity simulation of electron correlation and complex quantum effects, quantum algorithms offer a path to accuracy and scalability beyond classical limits, particularly for larger, intricate systems such as nanomaterials and metalloproteins. For routine property prediction, classical AI remains the state of the art, and quantum machine learning has no demonstrated advantage yet. The quantum opportunity is concentrated in the middle rows — binding prediction for metal-protein targets and high-accuracy energies for large molecules — where classical methods are weakest and the quantum mapping is most natural.
Real-World Impact and Future Outlook
Real-world signals are already appearing, though they remain proofs of principle rather than production results. St. Jude reports that Gorgulla's team identified ligands for one of the most important cancer drug targets and experimentally validated two molecules with real-world potential for future evaluation, calling the study proof-of-principle that quantum computing can greatly enhance drug discovery in the future. Drug Discovery World likewise points to IBM's collaboration with MIT researchers on calculating the ground-state energy of complex molecules, demonstrating that quantum computers could simulate electronic structure with an accuracy unattainable by classical methods — an achievement Turner reports paves the way for designing novel compounds in record time.
These milestones matter because they test the theory where it counts: not on toy molecules, but on targets where classical methods struggle. The Frontiers review's timeline gives the field its next dated markers — quantum-assisted QM/MM binding calculations remain ongoing research, while QPE for drug-sized molecules and Grover-based screening are placed after 2030. Until then, the honest picture is a hybrid one: DFT, FEP+, docking software and graph neural networks carry the daily workload of computational chemistry, while quantum methods advance on the problems those tools provably cannot reach.
Embracing Quantum for Intractable Molecular Challenges
The decision for researchers and pharmaceutical companies follows directly from the comparison: strategically investigate and adopt quantum computing for molecular simulation tasks that are currently intractable or highly inefficient for classical methods, particularly those involving complex electronic structure and electron correlation — while keeping DFT, FEP+ and graph neural networks in place for the tasks they still win. The practical entry points are the binding sites of metal-containing proteins, reactive intermediates and nanomaterials, where force-field approximations fail first and the quantum mapping is most direct.
The measurable indicator to watch is increased publication of quantum simulation results for novel drug candidates or materials properties, demonstrating superior accuracy or speed compared to classical benchmarks for specific complex systems. Head-to-head results of that kind — not theoretical speedups — are what will convert the shared-substrate argument into demonstrated advantage. Until those results appear at drug-relevant scale, the quantum advantage remains a mapped but unfinished frontier, with its next checkpoints already dated by the field's own review literature.